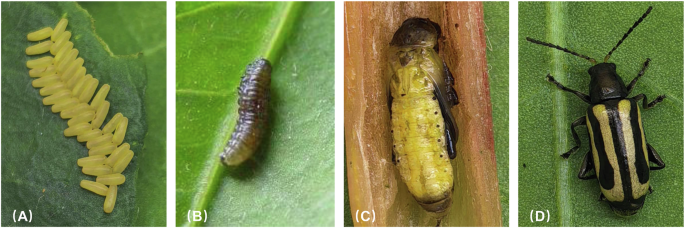
Researchers have completed a chromosome-level genome assembly for Agasicles hygrophila, a beetle used as a host-specific biocontrol agent, according to a study published in Scientific Data on Sunday.
The assembly provides a high-quality genetic map for the insect, which is widely utilized to manage invasive populations of A. philoxeroides.
To generate the data, scientists collected A. hygrophila larvae from Fuzhou, Fujian, China, and maintained the colony in a laboratory setting for approximately thirty generations. The sequencing process involved extracting high-molecular-weight genomic DNA from 5 grams of pooled third-instar larvae that had been flash-frozen in liquid nitrogen.
Researchers utilized three distinct library types to ensure assembly accuracy: short-read sequencing on the DNBseq platform, long-read HiFi sequencing on the PacBio Revio system, and Hi-C library construction to facilitate chromosomal scaffolding.
Initial genome surveys estimated the size of the beetle’s genome at approximately 716.98 Mb, characterized by a heterozygosity rate of 0.562% and a significant repeat rate of 63.3%. Following the initial de novo assembly, which produced 1,877 contigs, the team employed Hi-C data to order and orient these segments into chromosome-level scaffolds.
During this scaffolding phase, the researchers removed sequence segments that lacked consistent linkage, accounting for 11.19% of the initial assembly. The final chromosome-level genome measures 632.72 Mb and consists of 468 scaffolds.
The study reports that the short-read library produced approximately 95.33 Gb of clean data after quality filtering, while the PacBio HiFi library yielded 59.68 Gb of reads with an average length of 20,634 bp and an N50 of 20,920 bp. The Hi-C library generated 953,585,540 clean reads amounting to roughly 142.58 Gb of data, according to the authors.
DNA quality was assessed using a NanoDrop 2000 spectrophotometer and a Qubit 3.0 fluorometer, and integrity was evaluated by 0.5% agarose gel electrophoresis. High-molecular-weight DNA was extracted with a modified CTAB protocol from larvae that had undergone a 24-hour starvation period before flash-freezing.
De novo assembly was performed with NextDenovo v2.5.2 using HiFi data, followed by polishing with short reads via NextPolish v1.4.0.
Hi-C reads were processed with Juicer v1.6 and scaffolded using the 3D-DNA pipeline v180922, with manual review in Juicebox v1.11.08 and assistance from HapHiC v1.0.2. During scaffolding, contigs were split at misjoins and segments lacking consistent Hi-C linkage were removed, accounting for 79,711,006 bp (11.19% of the initial assembly), while 98,000 bp of N gaps were inserted between joined contigs.
The study, authored by Jian-Yu Li, Yan-Ting Chen, Jian-Wei Zhao, and their colleagues, establishes a comprehensive genomic resource for the species. By integrating short-read, long-read, and Hi-C technologies, the team successfully addressed the challenges posed by the high repeat content inherent in the A. hygrophila genome, ultimately achieving a final assembly where 90.03% of the sequence is anchored to chromosomes.
Worth a look